
Researchers at AstraZeneca in the UK have developed a computational workflow to predict if a compound will exhibit atropisomerism,1 an exotic form of chirality that can complicate drug development. The company has already incorporated the tool into its internal workflows.
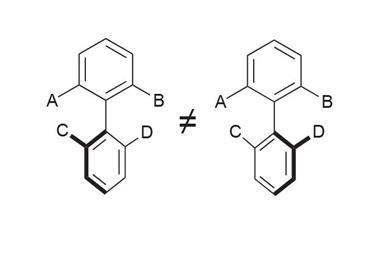
Atropisomerism is a type of axial chirality that occurs when bulky groups restrict rotation around a single bond.
Approximately 30% of recently approved FDA drugs have a potentially atropisomeric axis. Not all atropisomeric axes pose a problem for drug development; it depends on their energy barrier to rotation. In general, either a low or a high barrier is acceptable. A high barrier means rotation is slow and therefore unlikely to happen on a meaningful timescale; a low barrier means the molecule will rapidly interconvert, resulting in a racemic mixture.
The real problem is when the barrier to rotation sees isomers interconvert over timescales of minutes to months, meaning a molecule’s stereochemistry can change during storage or in the body, potentially affecting its potency, selectivity and safety profile. Such atropisomers have been described as a ‘lurking menace’ because they can remain hidden until late-stage development,2 when companies may find they have invested substantial resources in drug candidates that they later need to alter or abandon. ‘The error of landing on one of these unfortunate atropisomers may cost a lot of money to a pharmaceutical company,’ notes Art Bochevarov, a product manager at software company Schrödinger.
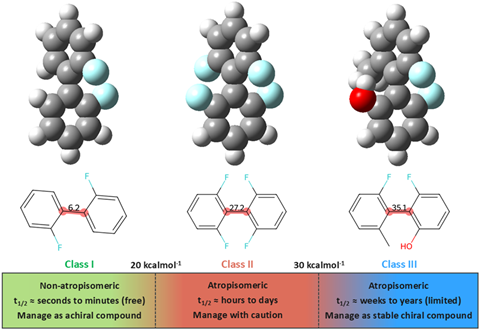
Led by Elliot Farrar, the AstraZeneca team’s tool uses cheminformatics and quantum mechanics in a modular fashion to assess molecules’ conformations and transition states under realistic solvent and temperature conditions, to pre-screen them for potentially troublesome atropisomeric axes. ‘There are already numerous internal projects at AstraZeneca where we’ve included rotational barriers calculated with our workflow,’ says Farrar. For example, the tool was recently used to aid the development of a lung cancer drug candidate.4
Earlier this year, Bochevarov and colleagues at Schrödinger reported their own computational tool for predicting atropisomerism in drug-like molecules.3 Both methods achieve similar levels of accuracy, but they differ in how they work.
AstraZeneca’s approach uses Smiles arbitrary target specification (Smarts) strings-based pattern matching to find restricted bonds including in complex systems. The Schrödinger tool searches each rotatable bond one at a time and can therefore find restricted bonds that may otherwise have been missed.
The tools also differ in accessibility. Schrödinger’s software comes with a user-friendly interface that makes it straightforward for non-specialists to use, but access requires a subscription. AstraZeneca’s tool, meanwhile, is openly available, but requires some computational experience to use it.
‘The biggest limitation that we share between both ours and the Bochevarov tool is addressing larger fused ring systems, particularly macrocycles,’ notes Farrar. Indeed, macrocycles are increasingly important in drug development, but offer a more complicated conformation landscape, which makes atropisomerism much harder to predict. ‘You cannot say which bond [in a macrocycle] is responsible for atropisomerism … essentially all of them are responsible,’ notes Bochevarov. Both groups are now working to solve this problem.
References
1 E H E Farrar et al, Digital Discovery, 2026, DOI: 10.1039/d6dd00210b
2 J Clayden et al, Angew. Chem., Int. Ed., 2009, 48, 6398 (DOI: 10.1002/anie.200901719)
3 Ty Balduf et al, J. Chem. Inf. Model., 2026, 66, 1675 (DOI: 10.1021/acs.jcim.5c02720)
4 J J Douglas et al, J. Org. Chem., 2022, 87, 2075 (DOI: 10.1021/acs.joc.1c01736)

No comments yet